MSE 298 Seminar: Challenges and Opportunities Using AI to Accelerate Materials Discovery and Process Optimization
Edward J. Schaefer Professor
Department of Chemical and Biomolecular Engineering
Johns Hopkins University
Abstract: The Materials Genome Initiative was started by NIST in 2011 with the prospect of using machine learning to make materials “faster, cheaper, better.” A decade or so later, we are finally close to having AI tools to exploit that prospect more effectively. A major factor contributing to the inefficiency of material discovery and design is the large combinatorial space of materials candidates as well as processing conditions, which are often sparsely observed for a given application. Searches of this space are often formed by expert knowledge and clustered close to known materials (exploitation). Exhaustive experimental characterization or first principles calculations are generally too expensive for an exhaustive search of this space. This invariably leads to small available data sets (<100 data points). As a result, there is a need to develop algorithms that can efficiently search this large parameter space and capable of dealing with “tiny” data sets. In this talk, we will introduce our approach to mitigate some of these issues using our in-house PAL2.0, Bayesian optimization codebase, that features a chemistry-informed belief model. A key characteristic of PAL 2.0 is the creation of a “physics-based” hypothesis (more often, chemistry-based) using a suitable feature selection tool and a simple Neural Networks as the front end to this process. This provides a generally informative physics-based prior (input) to the Gaussian process model used in Bayesian optimization. Our method picks out the physical descriptors that are most representative of the material domain, making the search independent from expert knowledge. Here, we demonstrate PAL2.0's use in challenging situations like polymer design using a computational, density functional theory, training set to predict high-performing conducting polymers. As a second test case, we show PAL’s versatility to use solely experimental data to create a “closed-loop” iterative loop of {predict-make-test-retrain} activities. This approach was performed with experimental collaborators at Hopkins Advanced Physics Laboratory to discover high-temperature shape memory alloys for space actuation applications. We use constraints to help “zero in” to manufacturable solutions in the fewest number of optimization iterations. As a result of this “closed loop” process, we discovered new multi-principal element alloys (MPEAs) with dramatically improved targeted properties, in this case, alloy toughness. More impressively, we find alloy candidates suggested by PAL2.0 that are “out of distribution,” which means that they suggested the use of elements (Si and Ta) that were not present in the initial database. PAL2.0 is able to “think out of the box.” We end with some of the current challenges in using AI for these tasks and attempt to deflate some of the hype.
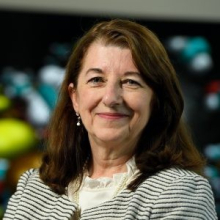
Bio: Paulette Clancy is the Edward J. Schaefer Professor of chemical and biomolecular engineering at the Johns Hopkins University. She is the Samuel and Diane Bodman Chair (emerita) at Cornell University. She was an inaugural director of research for Hopkins Data Science and AI Institute until July 2026. She has now moved to their Leadership and Strategy Council for the same AI Institute. She is Chair of the Research Computing Faculty Advisory Committee. She is a fellow of the Royal Society of Chemistry of AIChE and of the American Institute of Chemical Engineers. She spent over 30 years teaching and leading her department at Cornell for two terms before moving to Johns Hopkins in 2018 to become the inaugural department Head of ChemBE (2018-2023).
Paulette Clancy has one of the leading groups in the country studying atomic-scale modeling of semiconductor materials and sustainable energy applications. Her projects range from traditional silicon-based compounds to all-organic materials including flexible wearable organic electronics; electronic materials processing (III-V semiconducting materials and polymer-based thermoelectrics); and nucleation and crystal growth (hybrid organic/inorganic perovskites and quantum dot nanocrystals). Her lab focusses on studies of advanced materials processing to link processing, structure and function.
Over the past decade, her group has become best known for pioneering AI algorithm development for “materials discovery.” To that end, she developed new Bayesian optimization methods that encode chemistry and physics knowledge and intuition into the AI models. Her group is now expanding the term “materials discovery” to harness AI’s capability to help optimize materials processing protocols and not simply focus on predicting compositions that are silent on how these “materials” might be made. Recent machine learning projects involve “closed loop” AI-driven optimization of metal halide perovskites for solar cells, discovering new high-entropy alloys for spacecraft parts, and new Mg-alloys for orthopedic screw design. She is now turning to developing new AI methods for image processing and Machine-Learned Interatomic Potentials.
Share

Related Content
| Attachment | Size |
|---|---|
| 2.43 MB |
Upcoming Events
-
EECS Seminar: Printed Electronic Skin for Robots and Interactive Systems
-
MAE 298 Seminar:Ellipse Synthesis and the Computational Design of Robot Geometries
-
CBE 298 Seminar: Do I Belong Here? Addressing Imposter Syndrome in Graduate School
-
CEE Ph.D. Defense Announcement: Detection and Monitoring of Viruses in Water for Public Health & Environmental Engineering Applications
-
MSE 298 Seminar: Challenges and Opportunities Using AI to Accelerate Materials Discovery and Process Optimization